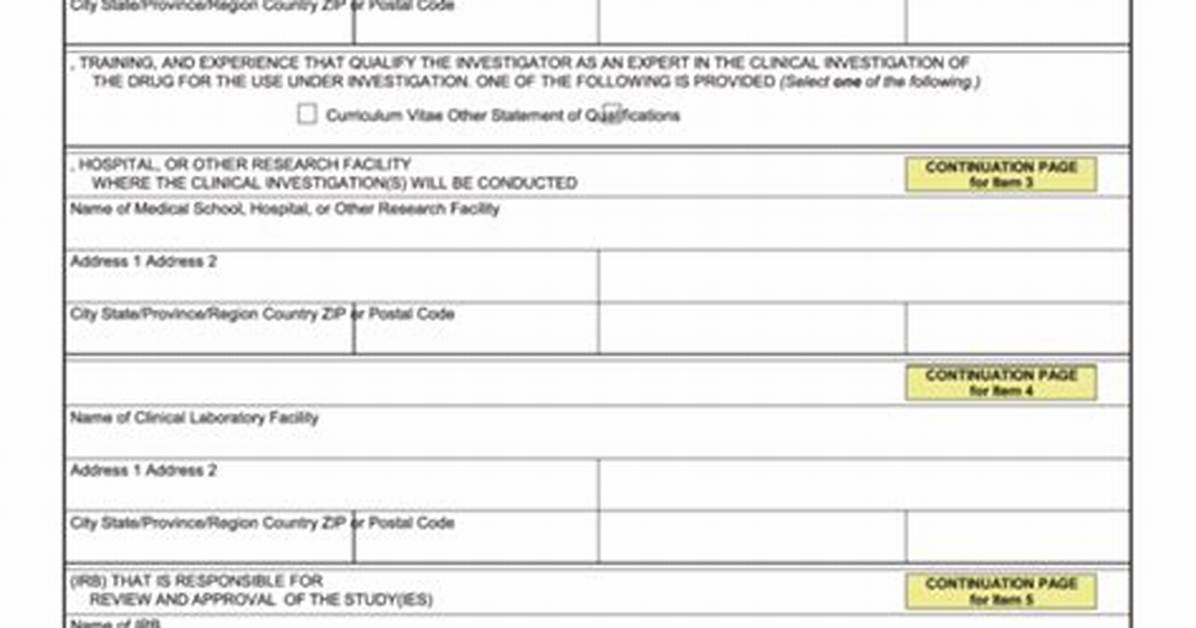
When Must the Form be Updated or a New One Completed? In cases when a new site is added or of replacement of an investigator at an existing site, a 1572 must be submitted to the FDA within a 30-day window of the site’s/investigator’s addition/replacement.
Who should be concerned by FDA Form 1572 and why?
Under the US regulations, all US clinical trial sites must be subject to the US IND and all US clinical trial investigators must sign Form 1572. All study sites (foreign or US) that are listed on the IND must comply with all applicable US regulations and the Principle Investigator (foreign or US) must sign Form 1572.
What is the purpose of FDA Form 1572 Mcq?
FDA Form 1572 is a legally binding document designed to inform clinical investigators of their research obligations and secure the investigators’ commitment to follow pertinent FDA regulations. By signing this form, the investigator confirms that they will abide by all FDA regulations.
Is a 1572 required for a device study?
Under FDA regulations, a 1572 is only required for studies of investigational drugs and biologics conducted under an IND. It is not required for studies that are not done under an IND and is not applicable to investigational device studies.
Is 1572 required IND?
The Draft Guidance highlights that clinical studies conducted at sites outside the U.S. under an IND are subject to all FDA IND regulations, including the requirement to obtain a signed Form 1572.
Is 1572 required for IND submission?
When filing an Initial IND Submission, a completed Form FDA 1572 must be sent from each site. When adding a new investigator (or new site), or replacing an investigator at an existing site. Note: a Form FDA 1572 must be submitted to the FDA within 30 days of the investigator being added.
Can study coordinators be on the 1572?
The FAQ states that study coordinators should”usually be listed in Section #6 of the 1572.” This is a significant change from the July 2008 draft FAQ. Many sites do not list studycoordinators on the FDA 1572, which means that most coordinators will need to file financial disclosure information.
What is a FDA Form 1571?
What is a FDA Form 1571? The FDA Form 1571 or ‘1571’ is the IND application cover page and it must accompany the initial IND submission and any amendments, IND safety reports, annual reports or general correspondence the sponsor submits to the FDA about the IND.
Who can report an adverse event in research?
If the investigator determines that the adverse event represents an unanticipated problem, the investigator must report it promptly to the IRB (45 CFR 46.103(b)(5)).
What is drug accountability?
What is means drug accountability? Drug accountability includes: study drug storage, handling, dispensing, and documentation of administration, return and/or destruction of the drug. A drug accountability process should be initiated for any study that uses study- supplied drug.
Who can be a sub investigator?
The ICH GCP 1.56 defines sub-investigator as any individual member of the clinical trial designated and supervised by the investigator at a trial site to perform critical trial-related procedures and/or make important trial-related decisions (e.g., associates, residents, research fellows).
What is an investigator’s commitment to the sponsor?
The sponsor shall obtain a commitment from the clinical investigator to promptly update this information if any relevant changes occur during the course of the investigation and for 1 year following the completion of the study.
What is an SAE in clinical trials?
SAE. SAE is short for Serious Adverse Event. An SAE is any untoward medical occurrence in a patient or trial subject, which does not have a causal relationship with the treatment, and: is fatal, and/or. is life-threatening for the subject, and/or.

